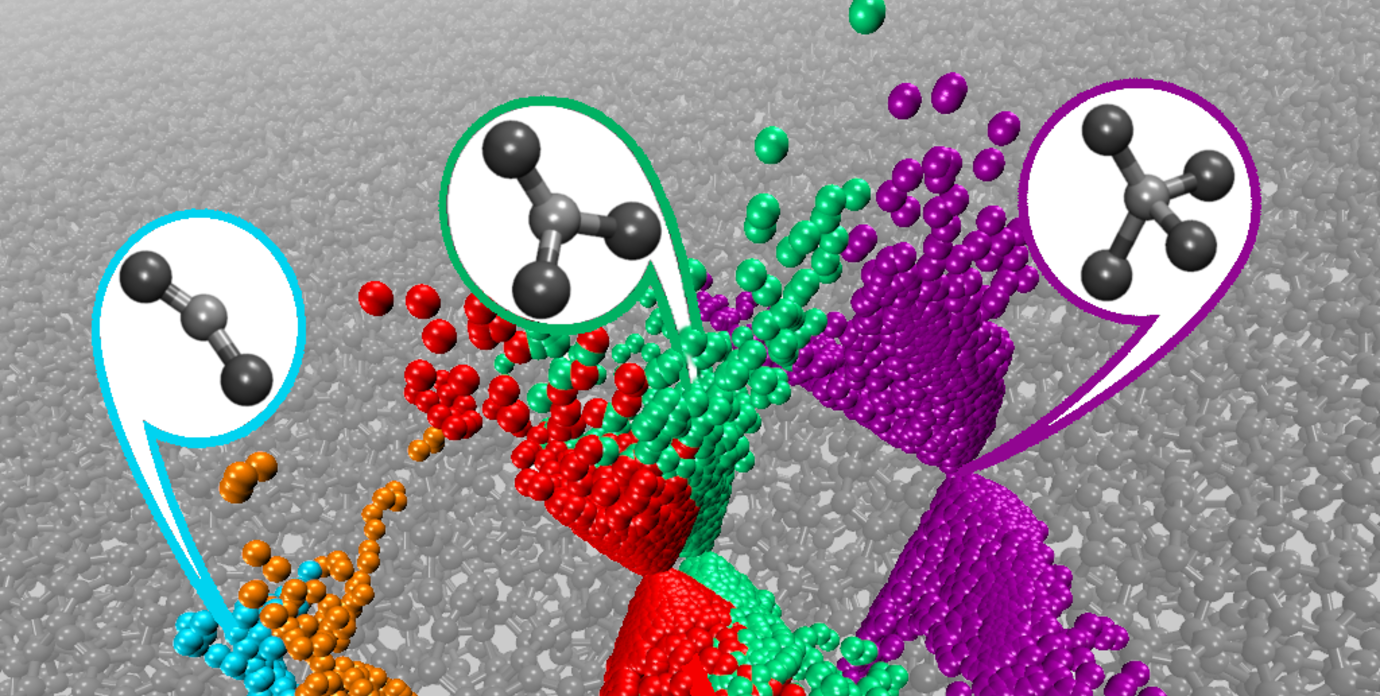
The infinite number of local atomic structures formed by carbon can be grouped into a few motifs with characteristic atomic and electronic properties.
The potential applications for tailor-made carbon surfaces are wide and include protective coatings, car parts, biomedical coatings and biosensors.
Yet for these developments to be realised, detailed atomic level knowledge is still needed on how carbon surfaces are structured and how they can be modified.
Thanks to the development of a new computational model, Postdoctoral Researcher Miguel Caro is spearheading work in this field by researchers at Aalto University, who work in partnership with Professor Gábor Csányi and Dr Volker Deringer from Cambridge University.
‘For the first time, we can identify the chemical properties of carbon surfaces and better understand how we can prepare them for specific purposes,’ explains Aalto University’s Professor Tomi Laurila.
The local environment of every atom in amorphous carbons, also called diamond-like carbons, is slightly different. This means that the number of neighbouring atoms, as well as the distances and angles between them, varies, posing a big challenge in the search to customize these surfaces.
The new computational model has finally allowed researchers to identify a wide variety of local atomic environments and classify them according to their properties. The research team has also calculated the varying degrees of strength with which different groups—hydrogen, alcohol (hydroxyl), and oxygen—will attach to surface sites.
Some bonds are, naturally, stronger than others. Because new information about the surface structures can be incorporated to ‘retrain’ and improve the model, the properties of still unknown surfaces can be predicted based on previous results.
‘Through computations, we can now not only explore what material surfaces look like at the atomic level but also see how they interact with other substances under analysis, as well as understand the kinds of chemical groups formed on these surfaces because of this interaction. We are also investigating what kinds of surfaces are needed to optimise the interaction with molecules that we would like to be able to detect, such as hydrogen peroxide,’ explains Laurila.
In other words, these simulation models based on density functional theory and machine learning tell us what kinds of structures can be developed—and how those structures may be optimised for specific applications.
‘In the future we will be able to produce tailored carbon surfaces, for example, for medical sensors, which could be used to monitor the concentration of a particular medication in a patient’s blood in real-time. Tracking changes in specific biomarkers in patients may be the key to improving therapeutic treatments currently used, or help us identify the risk of outbreaks of many common diseases earlier than ever before,’ Laurila says.
The study was published today as one of the cover articles in Chemistry of Materials.
Contacts:
Miguel Caro
Postdoctoral researcher, Aalto University
+358 50 407 9988
miguel.caro@aalto.fi
Tomi Laurila
Professor, Aalto University
+358 50 341 4375
tomi.laurila@aalto.fi
Dr Volker Deringer
Leverhulme Early Career Fellow, University of Cambridge
+44 7494 989967
vld24@cam.ac.uk
Aalto
#MaterialScience #Chemistry #ThinFilms #Nanostructures #Bionano #Sensors
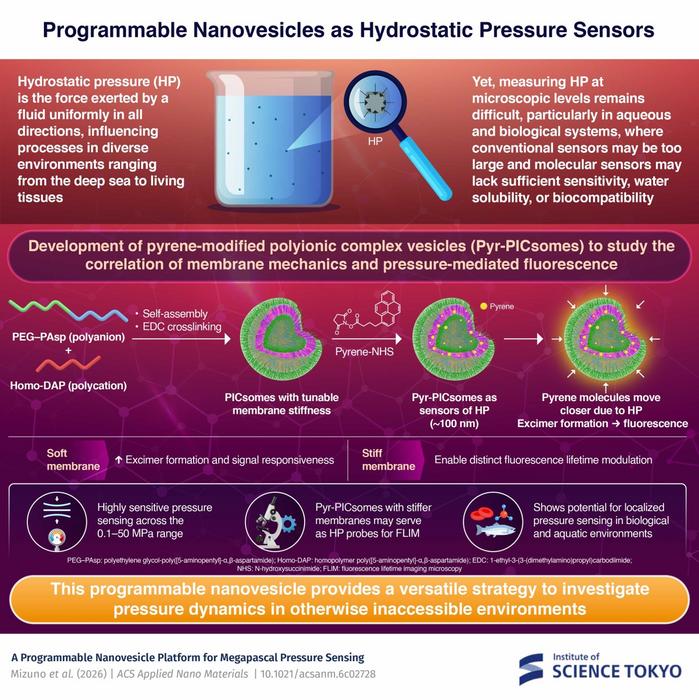
Pyrene-modified polyionic complex vesicles, or Pyr-PICsomes, convert hydrostatic pressure into measurable fluorescence. Crosslinking density tunes membrane stiffness, which governs sensing behavior: soft vesicles give strong excimer-to-monomer ratio changes from 0.1 to 50 MPa, while stiff membranes respond through fluorescence lifetime, suiting lifetime imaging.
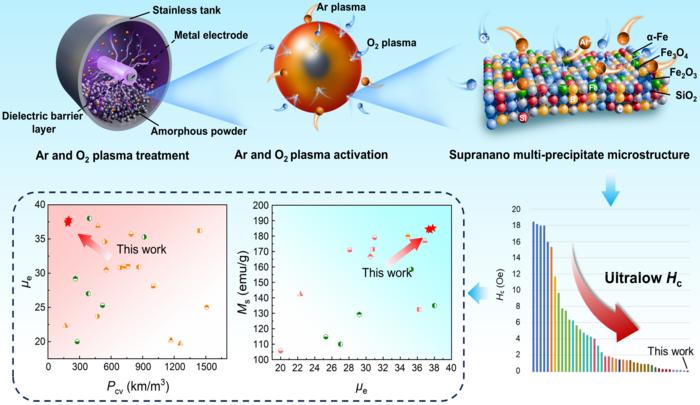
Ar/O₂ plasma treatment generates supranano α-Fe, Fe₃O₄, SiO₂ and Fe₂O₃ precipitates just 0.8–2.5 nm across amorphous powder surfaces, creating nucleation sites that ease magnetic-moment reversal. The resulting soft magnetic composite combines 0.13 Oe coercivity, 37.3 permeability, 185 emu g⁻¹ saturation magnetization and 191.18 kW m⁻³ core loss at 1 MHz and 20 mT. The surface route avoids bulk composition changes, aiding scale-up for high-frequency inductors and power chokes.
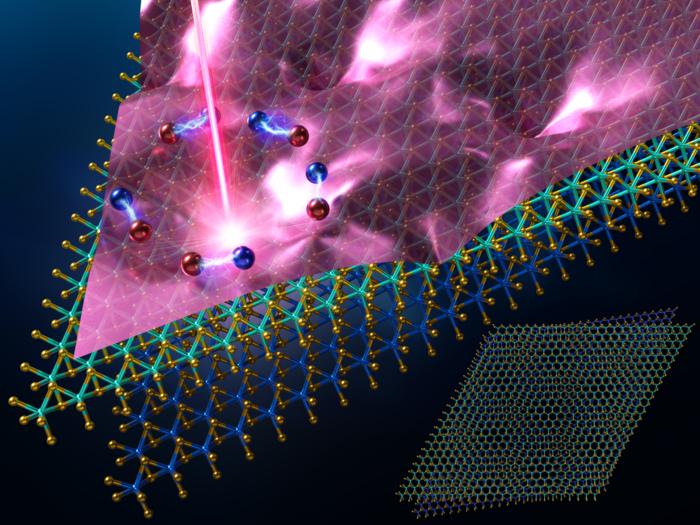
Researchers at MANA have introduced a theoretical framework that extracts hidden disorder from complex photoluminescence spectra in twisted MoSe2/WSe2 moiré heterostructures, bypassing unreliable peak-by-peak analysis. By tracking how simple descriptors like peak and average energy vary spatially, the work reveals a hierarchy of disorder, from micrometer-scale background variations to localized exciton-trapping defects.